TL;DR
We assess regulated-market API alternatives by the exact molecule and manufacturing site, not catalog size alone. This guide shows US formulation buyers how to compare containment, oncology, peptide and steroid fit, filing evidence, import readiness, development scale, and change control while clearly labeling evidence that is not publicly verified.
At the end of FY2025, FDA’s quality catalog included 5,953 manufacturing sites worldwide, a reminder that supplier qualification must reach the exact facility, not stop at a corporate name. The latest FDA quality report also recorded 1,248 drug-quality inspections that year.
We assess regulated-market API alternatives by matching the exact molecule and manufacturing site before comparing portfolios. For regulated markets, buyers should verify high-potency containment, development and commercial scale, current DMF or CEP evidence, inspection history, analytical controls, import readiness, and change control. No supplier is interchangeable across every API class.
We built this comparison framework for US formulation buyers evaluating complex small molecules, oncology APIs, HPAPIs, peptides, steroids, intermediates, and development-to-commercial support.
Which Capabilities Should Alternatives Actually Match?
We start with the requested drug substance, not a supplier’s widest category list. A credible comparison asks whether the same site can make the required API form, meet its impurity and particle-size expectations, support the needed annual volume, and provide documentation appropriate to the target market.
A product may appear in a public catalog while its DMF, commercial status, containment approach, and approved manufacturing site remain undisclosed. FDA explains that a Type II DMF’s initial completeness assessment does not replace the full scientific review needed to support an ANDA, which is why a listed filing alone is not enough. FDA DMF guidance
We use four evidence labels throughout our sourcing work:
- Comparable: The buyer has product-and-site evidence for the requested capability.
- Potentially Stronger: The evidence indicates a more specialized match for the requested need.
- Potentially Narrower: The supplier may fit one requirement but lacks demonstrated overlap elsewhere.
- Not Publicly Verified: Public information does not substantiate the specific product, site, or control being compared.
This approach prevents a broad portfolio from being mistaken for a broad fit. Buyers who need a wider market map can also review our API catalog comparison before beginning molecule-level diligence.
Which Regulated-Market API Alternatives Fit by Capability?
The closest fit changes with the molecule. A steroid formulation program may prioritize dedicated controls, impurity data, and US documentation. An oncology program may place containment, occupational exposure controls, and validated cleaning ahead of portfolio breadth. A peptide program needs separate evidence for synthesis, purification, characterization, and scale.
Complex Small Molecules and Oncology
For complex small molecules, we compare route-development support, process robustness, impurity strategy, solid-state knowledge, analytical transfer, and validation readiness. ICH Q11 places drug-substance manufacturing and development information in CTD Module 3, including the manufacturing process, control of materials, controls of critical steps, and process validation approach. ICH Q11
For oncology APIs, we do not assume every oncology supplier has equivalent experience with every potent class. The relevant questions are whether the requested molecule has been made at the proposed site, what containment regime applies, and whether commercial-scale process and quality evidence can be reviewed.
Steroid, Peptide, and Intermediate Fit
Steroid sourcing requires a specific check for process containment, cleaning validation, residual-risk controls, and the requested API form. Our steroid manufacturer comparison helps buyers distinguish a broad steroid listing from a qualification-ready supply option.
Peptide capability is distinct from conventional small-molecule API capability. Buyers should request details on synthesis strategy, purification, characterization, stability, scale range, and filing support for the exact peptide.
Capability Comparison Matrix
| Supplier Profile | Site Evidence | Molecule Classes | Oncology | HPAPI Containment | Peptides | Steroids | Development Stage | Commercial Scale | Filing Support | US Supply Evidence |
|---|---|---|---|---|---|---|---|---|---|---|
| Indian Steroid API Profile | Exact site required | Corticosteroids and hormones | Not publicly verified | Verify by product | Not publicly verified | Verify by product | Verify | Verify | Product-specific DMF or CEP required | Registration, listing, and entry evidence required |
| Indian Complex API Profile | Exact site required | Complex small molecules and intermediates | Verify by molecule | Verify by product | Not publicly verified | Verify by molecule | Verify | Verify | Product-specific filing evidence required | Product-and-site evidence required |
| Global HPAPI CDMO Profile | Exact site required | Potent and oncology APIs | Verify by molecule | Verify containment controls | Verify separately | Not publicly verified | Verify | Verify | Product-specific filing evidence required | Product-and-site evidence required |
| Peptide Specialist Profile | Exact site required | Peptide APIs | Verify by molecule | Verify where applicable | Verify by peptide | Not publicly verified | Verify | Verify | Product-specific filing evidence required | Product-and-site evidence required |
| Niche Chemistry Profile | Exact site required | Specialty APIs and intermediates | Verify by molecule | Verify where applicable | Not publicly verified | Verify by molecule | Verify | Verify | Product-specific filing evidence required | Product-and-site evidence required |
The matrix is deliberately conservative. We would rather mark a claim as not publicly verified than imply equivalence based on a therapeutic-area label.
How Should Buyers Verify HPAPI and Oncology Controls?
HPAPI qualification should test controls rather than marketing language. FDA’s Q7 guidance says dedicated production areas should be considered for highly pharmacologically active or toxic materials, including certain steroids and cytotoxic anticancer agents, unless validated inactivation or cleaning procedures are established and maintained. FDA Q7
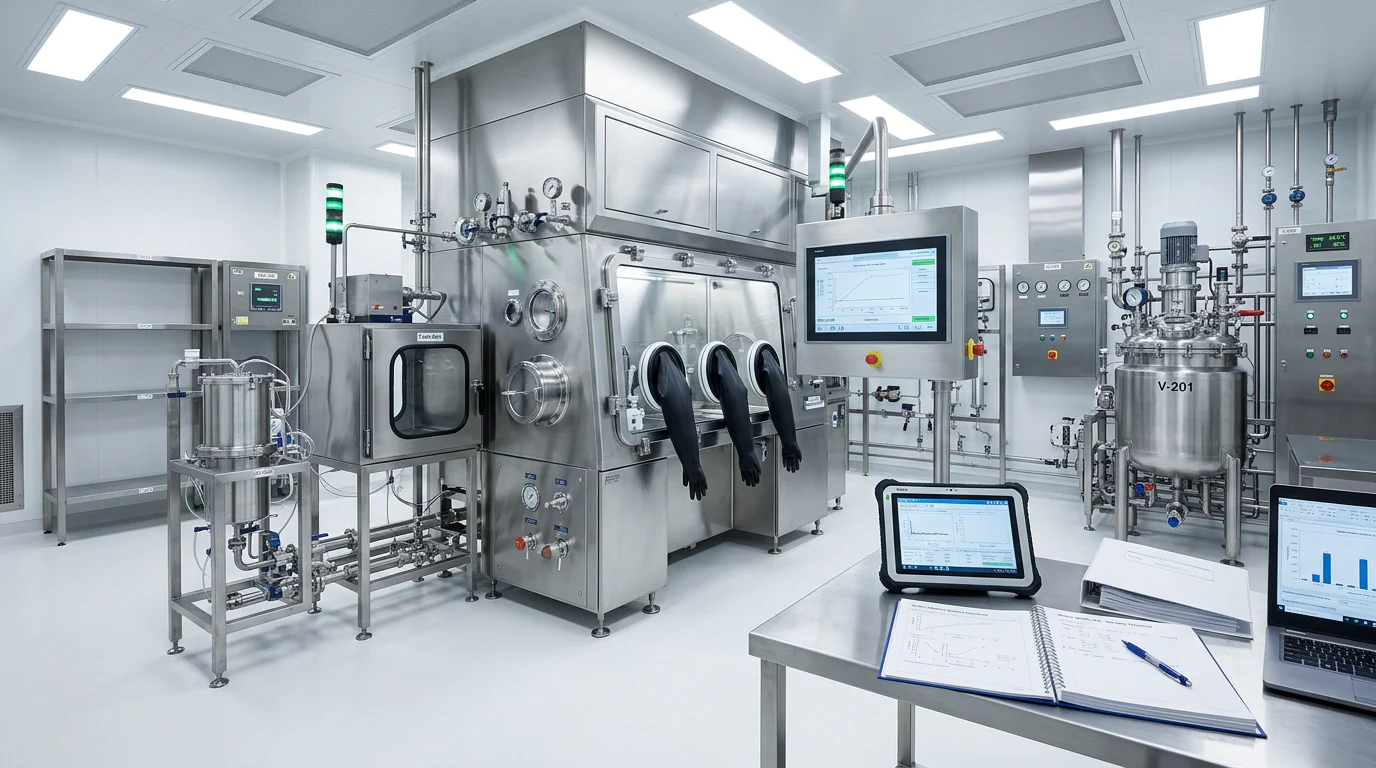
We ask for the evidence that connects a potent API to actual controls. That usually includes the occupational exposure rationale, containment design summary, material-transfer method, segregation rules, HVAC and pressure-cascade approach, cleaning-validation strategy, analytical detection capability, and deviation management process.
- Containment Design: Confirm whether the requested material is handled in closed equipment, isolators, or another documented containment system.
- Occupational Exposure: Request the exposure-band or limit rationale for the actual substance, not a generic potent-compound statement.
- Segregation: Check the flow of people, materials, waste, and equipment between potent and non-potent operations.
- Cleaning Validation: Review worst-case selection, residue acceptance criteria, sample locations, recovery studies, and analytical sensitivity.
- Change Control: Confirm how a new molecule, equipment train, starting material, or cleaning process is assessed before implementation.
FDA states that firms using non-dedicated equipment for potent compounds must define controls sufficient to prevent cross-contamination and validate cleaning procedures accordingly. FDA cleaning guidance Our GMP export guide provides the broader quality-system context buyers should bring into an audit.
Where Do Portfolios Overlap, and Where Do They Not?
Portfolio overlap is rarely all-or-nothing. We separate therapeutic labels from manufacturing reality because two suppliers can both list oncology, steroids, or peptides while supporting very different molecule types, scales, documentation packages, and controls.
| Buyer Need | Evidence That Supports A Strong Match | Evidence That Is Insufficient Alone |
|---|---|---|
| Complex Generic API | Process-development package, impurity strategy, solid-state support, validation plan, commercial batch history | General small-molecule catalog |
| Oncology Or HPAPI | Product-specific containment evidence, exposure rationale, cleaning validation, site qualification | Oncology category label |
| Peptide API | Synthesis, purification, characterization, stability, scale, and product-specific filing support | Research peptide availability |
| Steroid API | Exact form, potency controls, cleaning approach, analytical package, regulated-market documentation | Steroid or hormone listing |
| Intermediate Supply | Starting-material control, traceability, route transfer, and continuity plan | Ability to supply a related API |
Analytical capability deserves the same attention as process capability. We compare method transfer, reference standards, impurity profiling, residual solvents, elemental impurities, stability strategy, and data-review controls against the finished dosage form’s actual needs. Our quality testing guide explains the core testing questions that should accompany a request for quotation.
This is also where we distinguish development support from commercial support. A supplier may be valuable for route scouting or clinical material but not yet demonstrate the validated scale, capacity resilience, or change-control maturity a mature US generic program needs.
What Proves US and Other Regulated-Market Readiness?
A USDMF, CEP, ASMF, inspection record, and import history answer different questions. We do not treat any one of them as a substitute for the others.
USDMF and Letter of Authorization
A Type II DMF can support an API in a generic-drug submission, but the buyer still needs to confirm the DMF covers the exact drug substance, site, and intended reference. FDA’s DMF guidance specifies that a Letter of Authorization identifies the holder, DMF number, authorized party, covered product, submission details, and referenced sections. FDA LOA requirements
We recommend asking whether the DMF is current, active, available for reference when relevant, and aligned with the proposed manufacturing site. Our US filing guide gives buyers a practical companion checklist for that review.
CEP and ASMF Evidence
For European pathways, a valid CEP and an ASMF are not interchangeable with a USDMF. EMA explains that the ASMF procedure protects confidential manufacturing know-how while the marketing-authorisation holder remains responsible for the medicinal product’s quality and active-substance control. EMA ASMF guidance
We verify the substance, holder, site, certificate status, and timing. EDQM’s publicly accessible certification database provides current CEP status information, including valid, suspended, expired, and withdrawn certificates. EDQM database
Inspection and Import Evidence
Inspection evidence is site-specific and time-sensitive. FDA classifies closed inspections as No Action Indicated, Voluntary Action Indicated, or Official Action Indicated, while noting that not every inspection type appears in its public database. Inspection classifications
For US imports, foreign API manufacturers generally must register with FDA, list the API, and provide compliant import information. FDA compares declared manufacturer, importer or consignee, product description, intended use, and relevant affirmations against its records. FDA import requirements
How Should US Formulation Buyers Select a Final Supplier?
We recommend a six-step shortlist process that keeps technical, regulatory, and supply decisions connected. It is designed to identify the strongest regulated-market API alternatives for a specific formulation program, not to create a generic supplier ranking. For steroid programs, our US steroid import guide provides a useful companion for the import review.
- Define The Molecule: Specify API form, grade, critical quality attributes, forecast volume, dosage-form needs, target markets, and launch timing.
- Match The Site: Confirm where development, validation, commercial production, and quality control will occur.
- Verify Filing Evidence: Check the exact DMF, LOA readiness, CEP or ASMF status, and filing pathway for the requested product.
- Review Potent-Compound Controls: Assess containment, occupational exposure rationale, cleaning validation, segregation, and analytical sensitivity where applicable.
- Test Import Readiness: Confirm registration, listing, importer details, labeling expectations, documentation, and shipment data accuracy.
- Score Continuity And Change Control: Compare starting-material resilience, secondary options, lead times, quality agreements, notification commitments, and post-approval change governance.
ICH Q12 emphasizes that effective pharmaceutical quality systems must manage manufacturing changes across the supply chain, particularly when established conditions may require reporting to regulators. ICH Q12
The right choice is the supplier that can document the requested molecule, site, controls, filing support, and continuity plan most clearly.
Work with Aarise Pharma
At Aarise Pharma, we start with the evidence that keeps a qualification moving: the exact molecule, dosage-form needs, anticipated annual volume, target filing pathway, and required documentation. We help teams organize a practical technical request, compare supplier-provided records with regulator databases, and identify unresolved questions before a sample, audit, or commercial commitment. Our conversations focus on molecule-and-site verification, analytical expectations, change control, import readiness, and contingency planning. That gives formulators a clearer starting point for qualifying a steroid, peptide, oncology, or complex API, without pretending every supplier is interchangeable. We also help align development, validation, commercial scale, shipping, and post-approval scenarios with the questions quality units ask first. Bring us your specification, target market, and critical quality requirements, and we will help frame the next diligence questions. Talk with Aarise Pharma
FAQs on Regulated-market API Alternatives
What Makes an API Supplier Comparable?
Comparable suppliers match the requested molecule, manufacturing site, critical quality attributes, containment controls, development stage, and filing evidence. Broad therapeutic claims cannot replace product-and-site proof.
Does a USDMF Mean FDA Approval?
No. A Type II DMF may be available for reference, but FDA separately evaluates the supporting application, exact product, manufacturing site, and underlying scientific data.
What HPAPI Evidence Should Buyers Request?
Buyers should request exposure rationale, containment design, segregation controls, cleaning-validation records, analytical sensitivity, deviation management, and evidence that applies directly to the requested molecule and site.
How Can Buyers Verify Steroid API Supply for the United States?
For steroid programs, verify API form, site, documentation, registration, listing, import-entry accuracy, cleaning controls, analytical package, and import requirements before qualification with your quality team.
What Is the Difference Between a CEP and an ASMF?
A CEP demonstrates European Pharmacopoeia suitability for an active substance, while an ASMF protects confidential manufacturing information that supports a medicinal-product application held by another party.
Why Should Comparisons Be Reviewed Quarterly?
DMF statuses, CEP statuses, inspection findings, registrations, listings, capacity, supply risks, and product portfolios can change. Quarterly review keeps comparisons tied to current, product-specific evidence.