TL;DR
Score program stage, formulation complexity, transfer risk, accountability, and supply resilience. Choose an integrated partner when unresolved API and drug-product decisions need one coordinated team. Choose specialists when a dosage-form or peptide capability is the decisive need. In both models, name every site, set transfer criteria, and assign quality responsibilities.
Score five gates before choosing one peptide API and drug-product partner: program stage, formulation complexity, transfer risk, accountability, and supply resilience. Choose integration when unresolved API and product decisions need one coordinated team. Choose separate specialists when a specific peptide or dosage-form capability is the decisive need.
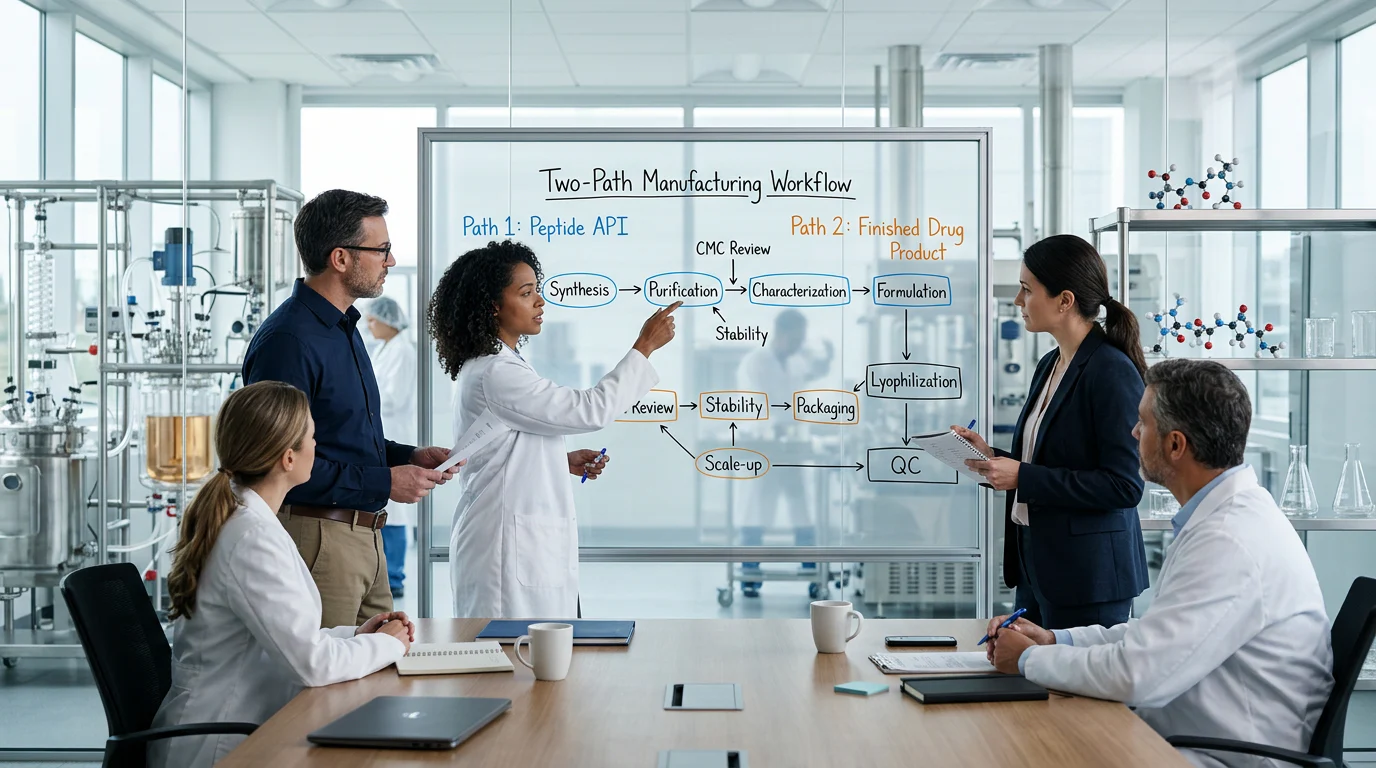
A peptide API is the active pharmaceutical ingredient, also called drug substance. Drug product is the finished dosage form, such as an injectable, tablet, or capsule. The sponsor remains responsible for controlling outsourced work under ICH Q10, whichever model it chooses.
Choose the Model with Five Gates
Use this table before sending an RFI. It turns a vague “one partner or two?” question into evidence each candidate must provide.
| Decision Factor | Integrated Path | Specialist Partners | Request |
|---|---|---|---|
| Program stage | API and product choices open | Mature API, independent product work | Development plan, named sites |
| Formulation complexity | Proven API-to-dose-form capability | Required dosage-form specialist | Stability and method plans |
| Transfer risk | Shared API and product teams | Complete transfer package ready | Protocol, acceptance criteria |
| Accountability | One RACI across every site | Sponsor coordinates both partners | Quality agreement |
| Supply resilience | Qualified alternate route exists | Qualified second source exists | Materials and capacity map |
The table points to a starting model, not a purchase order. Our rule of thumb: choose the model that wins three gates, unless the other model solves a required technical capability that the first cannot show.
When One Integrated Partner Reduces Risk
An integrated model fits best when the API and finished product are still shaping each other. That often happens when the team is deciding the API’s salt or counterion, solubility target, impurity limits, storage conditions, assay basis, or physical form.
ICH Q11 connects drug-substance development to drug-product needs. It specifically notes that drug-substance properties, including solubility, can affect dosage-form selection.
Choose one integrated partner when all three conditions apply:
- The peptide API process needs development, not only manufacture.
- The drug product needs formulation work that depends on API properties.
- The proposed sites can show the exact product capability your program needs.
A shared corporate name does not replace a transfer plan. Ask which legal entity and site will synthesize, purify, test, formulate, manufacture, release, and store your material. ICH Q10 notes that pharmaceutical quality-system effectiveness is normally demonstrated at the site level. Read ICH Q10
Integration also helps smaller sponsor teams. One project structure can reduce duplicate meetings and make one group responsible for the API-to-product handoff. The quality agreement should still identify each site, each subcontractor, and each escalation path.
When Separate Peptide Specialists Fit Better
Separate partners fit best when one stage needs depth that the other partner cannot demonstrate. A peptide synthesis specialist may be the stronger choice for a difficult sequence, purification challenge, impurity profile, or scale-up route. A different drug-product specialist may be stronger for a demanding formulation or finished-dosage process.
This choice creates an external technology transfer. Technology transfer means moving the process, methods, materials, and product knowledge from one organization to another so the receiving team can reproduce controlled results.
For EU-facing synthetic peptide programs, the current EMA peptide guideline makes the interface especially important. It calls for formulation work to address aggregation where relevant and applies peptide-related impurity thresholds to finished products: report above 0.1%, identify above 0.5%, and qualify above 1.0%.
Choose separate partners when these facts are true:
- The API route is sufficiently mature for another site to reproduce.
- A specialist has proven experience with the required product type.
- Your CMC, quality, regulatory, and supply teams can govern two partners.
- Both partners accept the same transfer criteria before work starts.
Separate partners can also support supply resilience. That benefit only exists when the sponsor develops an executable transfer route. Two suppliers with no shared method package, no material samples, and no comparability plan are two points of failure.
What Must Move from API to Drug Product
The transfer package should let the drug-product team understand the material, test it, store it, formulate it, and investigate a problem. Send the package before the first formulation batch, not after an out-of-specification result.
Include these technical records:
- Identity and form: sequence, modifications, salt or counterion, physical form, and packaging configuration.
- Quality attributes: assay, peptide-related impurities, process impurities, water, residual solvents, elemental impurities, and microbiological limits where relevant.
- Analytical methods: method purpose, system suitability, reference standards, sample preparation, acceptance criteria, and method-transfer responsibilities.
- Material handling: storage range, shipping conditions, hold times, retest or expiry basis, and excursion-response process.
- Process knowledge: synthesis route summary, purification approach, known degradation pathways, expected yield range, and critical process parameters.
- Product-development inputs: dose concentration, excipient compatibility questions, container-closure assumptions, and stability-indicating method status.
FDA’s API GMP guidance requires controlled records, including development, scale-up, technical-transfer, and validation reports. The same guidance requires written agreements that define GMP and quality responsibilities between contractors.
Do not equate an HPLC area percentage with usable peptide mass. Water, counterions, residual solvents, and assay basis can change the amount of active peptide delivered to formulation. Use our guide to separate HPLC purity from peptide content before comparing lots.
Set Accountability Before the First Batch
The quality agreement should answer who owns every decision at the interface. FDA describes quality agreements as the tool that delineates CGMP manufacturing activities between parties. FDA quality agreements
Assign these responsibilities by name:
- Approving API and drug-product specifications.
- Releasing API for formulation use.
- Approving analytical-method transfers.
- Investigating deviations, OOS results, and stability trends.
- Approving changes to materials, process, methods, sites, and specifications.
- Maintaining reference standards and retention samples.
- Notifying the sponsor of supply interruptions.
- Authorizing subcontractors.
- Escalating a quality issue across both workstreams.
A useful RACI lists one person as Responsible, one as Accountable, consulted technical groups, and informed stakeholders. Use one RACI across all sites, even if a single CDMO owns both operations.
For stage-specific deliverables, use our peptide CDMO process-development guide.
Protect Supply If One Partner Owns Both Steps
An integrated relationship can simplify operations while concentrating risk. Protect continuity by treating API manufacture and drug-product manufacture as separate operational nodes, even when one organization owns both.
Start with these actions:
- Name the primary manufacturing site for each step.
- Map every critical starting material, resin, reagent, excipient, and packaging component.
- Reserve capacity by campaign window, batch size, and release timing.
- Set notification periods for site, process, method, and material changes.
- Secure access to batch records, methods, reference standards, and stability data.
- Define the conditions for transferring work to another site or partner.
- Start second-source feasibility before a missed campaign threatens launch supply.
ICH Q10 expects companies to qualify outsourced activities, define responsibilities in written agreements, monitor contractor performance, and control materials through the approved supply chain. See outsourced-activity controls
A second source is not a second quotation. The incoming partner needs product understanding, material samples, analytical methods, comparability criteria, and enough process knowledge to make acceptable API or product. Use our peptide API supply-chain checklist to assign those workstreams.
FAQs
Does One CDMO Contract Mean One Manufacturing Site?
No. One contract can cover several legal entities, facilities, laboratories, and subcontractors. Ask for a site map that identifies where each activity occurs and who releases each intermediate or finished batch.
Is a 99% HPLC Result the Same as 99% Active Peptide?
No. HPLC area percentage measures a chromatographic signal under a stated method. The usable peptide amount can also depend on assay, water, counterion, residual solvents, and sample basis.
How Should We Compare Two Similar CDMO Quotes?
Compare each quote against one written scope. Require both candidates to name the sites, development outputs, transfer records, analytical work, stability plan, batch release path, change-control process, and continuity commitments included in the price.
Who Should Attend the Final CDMO Selection Meeting?
Include CMC, drug-product development, quality assurance, regulatory affairs, clinical operations, procurement, and supply planning. Each group owns evidence that the others cannot replace, from formulation risk to quality agreements and campaign timing.
At Aarise Pharma, we help pharmaceutical buyers structure peptide API requirements around the exact molecule, market, quantity, and documentation needed for a controlled supply decision. Explore our peptide CDMO stage guide before sending your next RFI.